目標基因
R 中的 Bioconductor 入門

Paula Andrea Martinez, PhD.
Data Scientist
基因組區間範例
- 對齊到參考序列的 reads
- 目標基因
- 外顯子區域
- 單核苷酸多型性(SNP)
- 轉錄區或結合位點、RNA-seq 或 ChIP-seq
Genomic Ranges
library(GenomicRanges)
(myGR <- GRanges("chr1:200-300"))
 GRanges object with 1 range and 0 metadata columns:
  seqnames     ranges strand
     <Rle>  <IRanges>  <Rle>
[1]     chr1 [200, 300]    *
 -----
seqinfo: 1 sequence from an unspecified genome; no seqlengths
- GRanges 類別用來依染色體儲存基因組區間
- 最少引數
chr1:200-300 - GRanges 的
seqnames與seqinfo
# df 類似 data.frame 的結構
   seqnames start end strand score  GC
1     chrX    50 120      +     1 0.25
2     chrX   130 140      +     2 0.25
3     chrX   153 154      +     3 0.25
4     chrY    30  40      *     4 0.25
5     chrY    50  55      -     5 0.25
(myGR <- as(df, "GRanges")) # 將 df 轉成 GRanges
GRanges object with 5 ranges and 2 metadata columns:
      seqnames     ranges strand |     score        GC
         <Rle>  <IRanges>  <Rle> | <integer> <numeric>
  [1]     chrX [ 50, 120]      + |         1      0.25
  [2]     chrX [130, 140]      + |         2      0.25
  [3]     chrX [153, 154]      + |         3      0.25
  [4]     chrY [ 30,  40]      * |         4      0.25
  [5]     chrY [ 50,  55]      - |         5      0.25
 -----
 seqinfo: 2 sequences from an unspecified genome; no seqlengths
Genomic Ranges 存取器
methods(class = "GRanges") # 檢視可用的存取器# 染色體名稱 seqnames(gr)# 回傳區間的 IRanges 物件 ranges(gr)# 儲存中繼資料欄 mcols(gr)# 通用函式,儲存序列資訊 seqinfo(gr)# 儲存基因組名稱 genome(gr)
- 存取器同時可設值也可取值
- 存取器可透過 S4 定義被繼承
目標基因:ABCD1
- ABCD1 位於 X 染色體長臂末端
- 在哺乳類的大腦與肺細胞功能中,負責編碼重要蛋白
- chrX 約 156 百萬 bp
- 座落於 chrX 約 153.70 百萬 bp
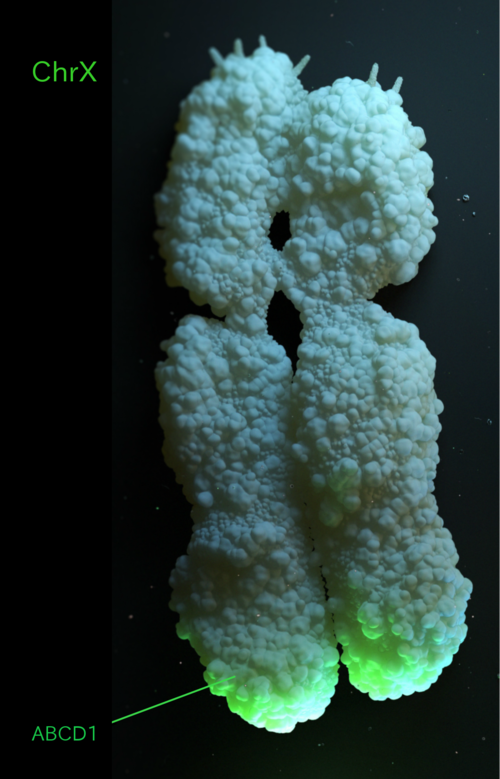
X 染色體的 GRanges
library(TxDb.Hsapiens.UCSC.hg38.knownGene)
hg <- TxDb.Hsapiens.UCSC.hg38.knownGene
從 X 染色體選取基因
hg_chrXg <- genes(hg, filter = list(tx_chrom = c("chrX")))
GRanges object with 1192 ranges and 1 metadata column:
            seqnames              ranges strand |     gene_id
               <Rle>           <IRanges>  <Rle> | <character>
  100008586     chrX   49551278-49568218      + |   100008586
      10009     chrX 120250752-120258398      + |       10009
  100093698     chrX   13310652-13319933      + |   100093698
        ...      ...                 ...    ... .         ...
  -------
  seqinfo: 640 sequences (1 circular) from hg38 genome
一起來練習吧!
R 中的 Bioconductor 入門

