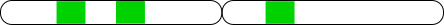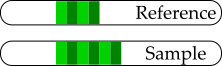清理 ChIP-seq 資料

在 R 中使用 Bioconductor 進行 ChIP-seq

Peter Humburg

Statistician, Macquarie University

常見問題

錯誤比對的 reads 會產生假峰值。

  • 基因體重複序列。

在 R 中使用 Bioconductor 進行 ChIP-seq

常見問題

錯誤比對的 reads 會產生假峰值。

  • 基因體重複序列。

  • 參考序列不完整。

在 R 中使用 Bioconductor 進行 ChIP-seq

常見問題

錯誤比對的 reads 會產生假峰值。

  • 基因體重複序列。

  • 參考序列不完整。

  • 低複雜度區域。

在 R 中使用 Bioconductor 進行 ChIP-seq

在 R 中使用 Bioconductor 進行 ChIP-seq

擴增偏差

  • 從細胞萃取的 DNA 片段在定序前會被多次擴增。
  • 並非每個片段都產生相同的複本數。
  • 相同片段的多個複本可能被定序。
  • 單一 DNA 片段會放大覆蓋度,導致錯誤的峰值判定。
在 R 中使用 Bioconductor 進行 ChIP-seq

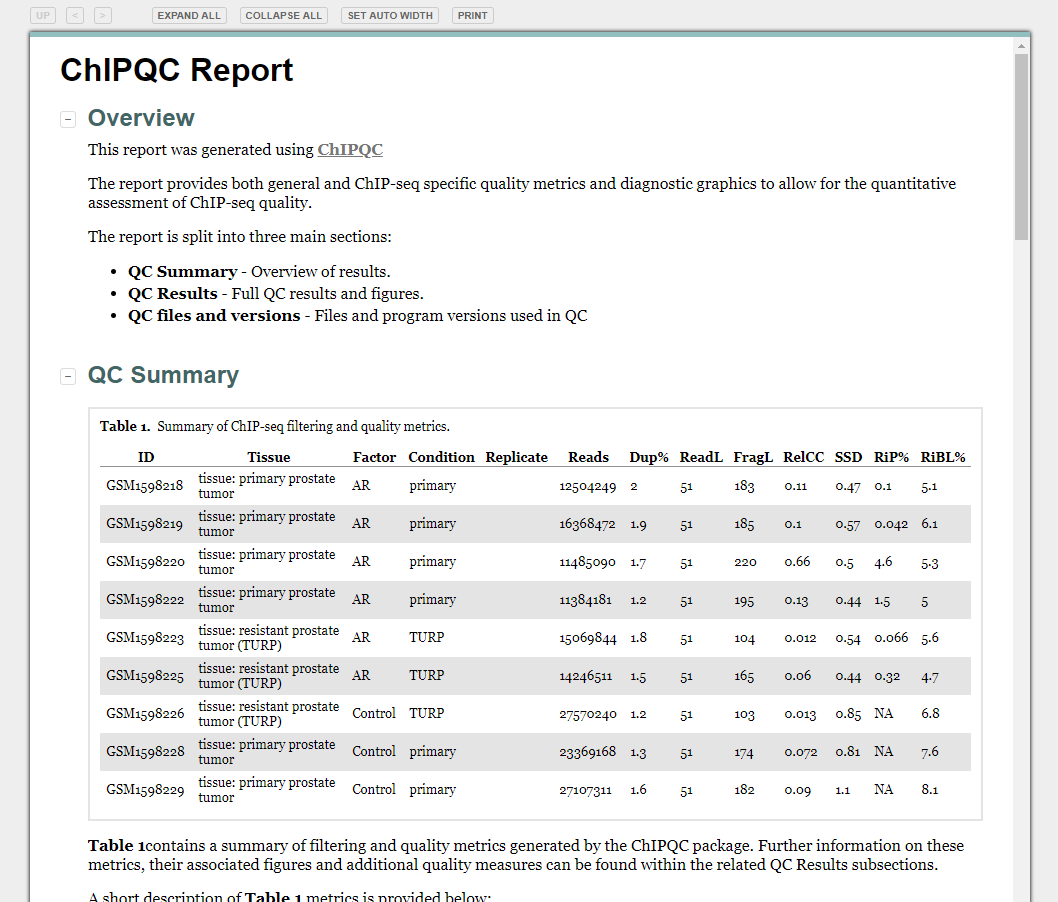
品質控制報告

library(ChIPQC)
qc_report <- ChIPQC(experiment="sample_info.csv", annotation="hg19")
ChIPQCreport(qc_report)
在 R 中使用 Bioconductor 進行 ChIP-seq

準備輸入檔

SampleID Factor Condition Tissue Treatment bamReads Peaks PeakCaller
S1 AR primary primary prostate tumor gleason score: 3+4=7 S1.bam S1.bed macs
S2 AR primary primary prostate tumor gleason score: 3+4=7 S2.bam S2.bed macs
... ... ... ... ... ... ... ...
在 R 中使用 Bioconductor 進行 ChIP-seq

清理資料

  • 移除重複 reads。
  • 移除多重比對的 reads。
  • 移除比對品質低的 reads。
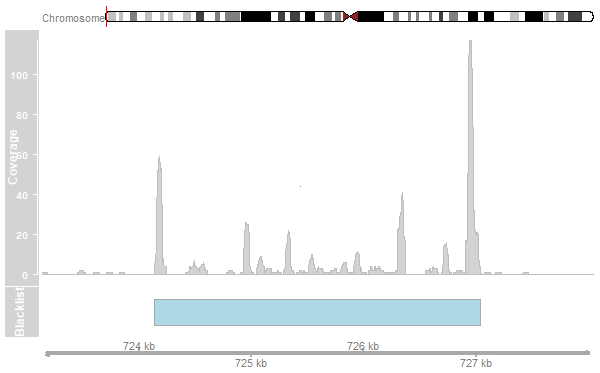
  • 移除黑名單區域的峰值。
在 R 中使用 Bioconductor 進行 ChIP-seq

清理資料

  • 移除重複 reads。
  • 移除多重比對的 reads。
  • 移除比對品質低的 reads。
  • 移除黑名單區域的峰值。
在 R 中使用 Bioconductor 進行 ChIP-seq

清理資料

  • 移除重複 reads。
  • 移除多重比對的 reads。
  • 移除比對品質低的 reads。
  • 移除黑名單區域的峰值。
    • 黑名單區域可從 ENCODE 計畫取得。
在 R 中使用 Bioconductor 進行 ChIP-seq

一起來練習吧!

在 R 中使用 Bioconductor 進行 ChIP-seq

Preparing Video For Download...